


OMIM ID:
Pigmented Paravenous Chorioretinal Atrophy
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
This is a rare type of pigmentary retinopathy with few symptoms in many patients. Pigment clumps in the form of bone spicules in a paravenous distribution appear as young as 1 year of age and may be present congenitally. The pigment may begin peripherally and is often segmental but eventually progresses centrally along with chorioretinal atrophy involving the majority of the fundus. For unknown reasons, males are more severely affected than females. In one family the retinal changes were associated with hyperopia, esotropia and vitreous degeneration (cells and liquefaction). There is considerable variation in expressivity among patients and the vision and fundus pigmentation can be highly asymmetrical in the two eyes. ERG abnormalities likewise vary widely with decreased photopic responses in some individuals and complete lack of both scotopic and photopic responses in severely affected eyes. Decreased night vision is not a symptom.
This is generally considered to be a stationary condition but long term follow up reveals progression of pigmentary changes, chorioretinal atrophy and increasing constriction of the peripheral visual field. Symptoms of decreased vision may be noted as early as 3 months of age. Some patients retain vision of 20/20 or 20/30 into midlife whereas others in the first decade already have count fingers vision. Likewise the size of the visual field varies widely and is not correlated with age.
Systemic Features
No systemic abnormalities have been reported.
Genetics
Inheritance
This is an autosomal dominant disorder caused by heterozygous mutations in the crumbs homolog 1 (CRB1) gene (1q31.3).
CRB1 mutations have been identified in other retinal disorders including nanophthalmos with retinitis pigmentosa, pigmented paravenous chorioretinal atrophy (172870), retinitis pigmentosa-12 (600105), and Leber congenital amaurosis 8 (613835). No consistent retinal phenotype has been found, however. There is often marked asymmetry between the two eyes and the rate of visual loss varies widely. Most individuals have some patchy areas of hypoautofluorescence in the posterior pole with variable amounts of pigmentary anomalies from mild speckling to frank bone spicule formation.
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
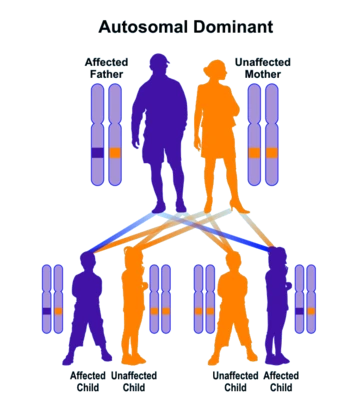
Autosomal dominant inheritance leads to a vertical pattern of transmission